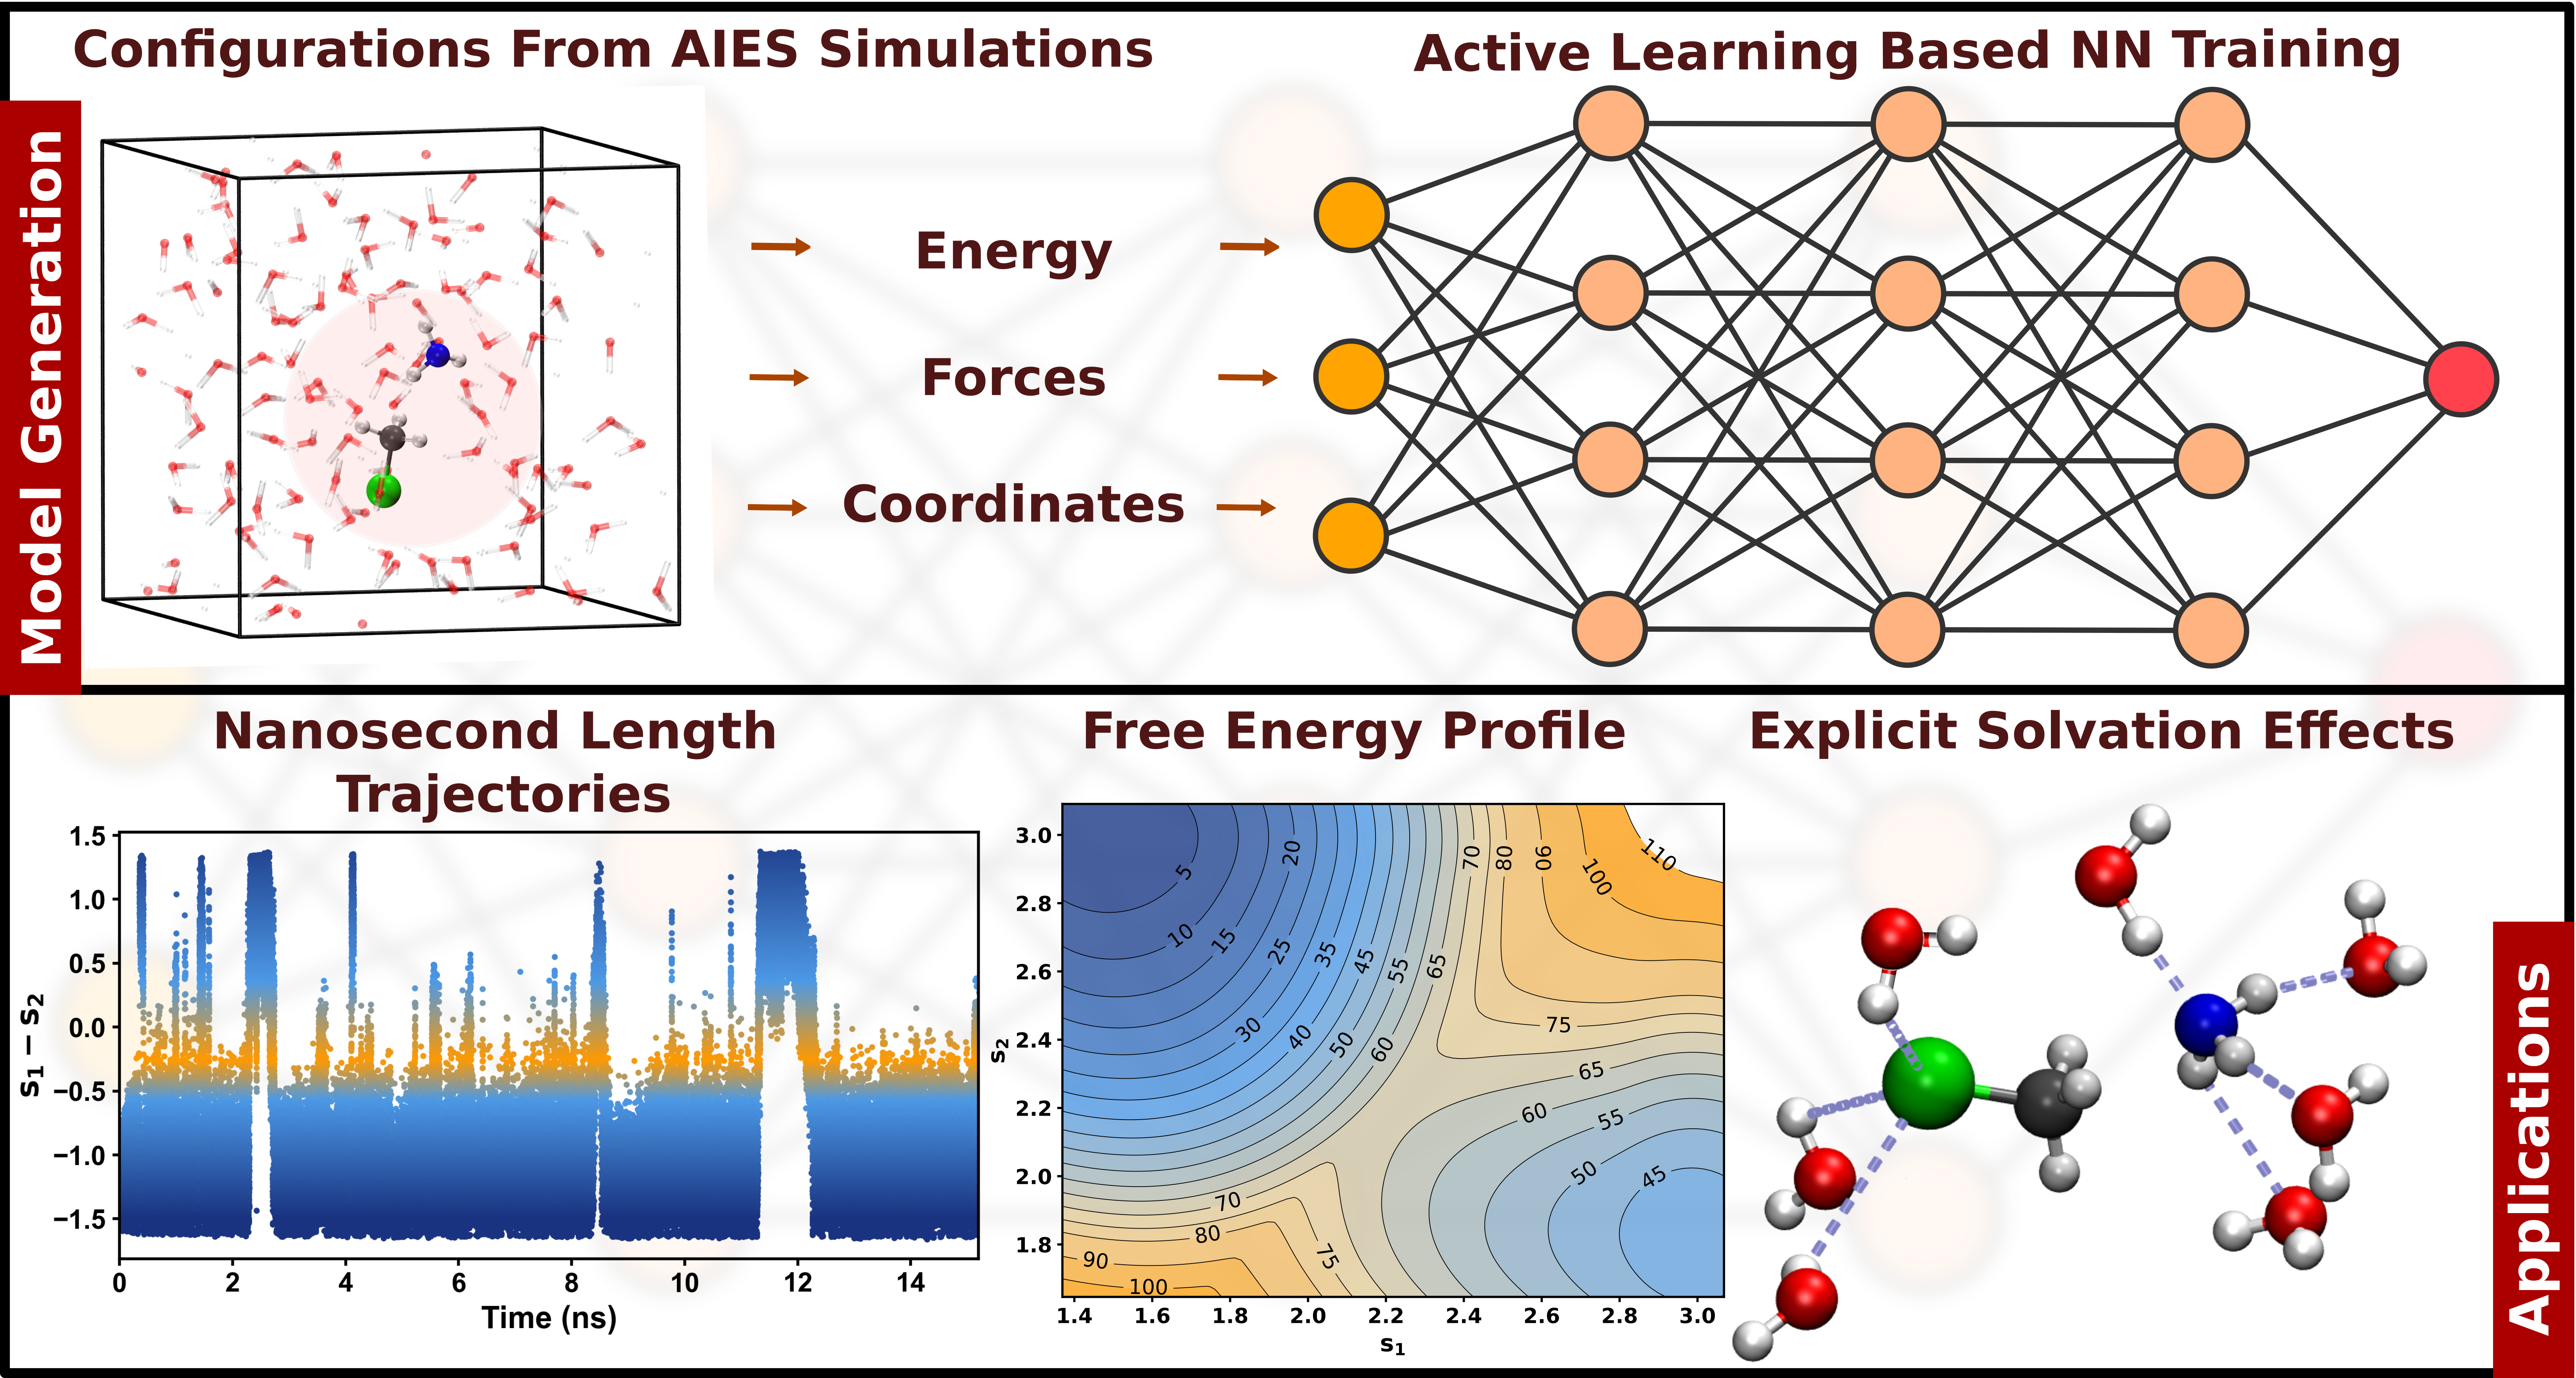
Chemical Reaction Modeling using ML-Augmented Enhanced Sampling
Developing advanced computational frameworks that combine machine learning with enhanced sampling techniques to understand complex chemical reaction mechanisms at the molecular level.
Exploring the frontiers of computational chemistry through machine learning potentials, enhanced sampling simulations, and quantum mechanical methods.
I am a research scholar in Computational Chemistry at the Indian Institute of Technology Delhi, originally from Hisar, Haryana. Under the supervision of Prof. Tarak Karmakar, my doctoral research focuses on the atomistic modeling of condensed-phase systems integrating quantum chemistry, molecular dynamics, and machine learning approaches.
I employ a wide range of computational methods including DFT, ab initio molecular dynamics, and enhanced sampling techniques such as metadynamics to explore free energy surfaces and simulate rare events. I also develop and apply machine learning interatomic potentials to scale up simulations of reactive systems while retaining quantum accuracy.
Through high-performance computing and advanced simulation techniques, my work bridges the gap between theoretical models and experimental observations, providing molecular-level insights that are challenging to obtain through experimental methods alone.
Developing advanced computational frameworks that combine machine learning with enhanced sampling techniques to understand complex chemical reaction mechanisms at the molecular level.

Creating innovative machine learning approaches to develop accurate interatomic potentials with minimal training data, enabling large-scale simulations with quantum mechanical accuracy.